What's new in olomol
The changes in each version, newest on top.
0.5.1 20 November 2026 newest
The manual and quick start for 0.5.0, and the manual as a page on the web too.
Changed
- The manual is now also a page on olomol.org, in English and Czech, with its contents beside the text: Manual in Help opens it, and PDF beside it is the manual for printing. The search bar finds both.
- The manual and the quick start describe 0.5.0: tabs and Show, .olom files and a drawing’s earlier versions, the search bar, groups and the Objects panel, drawing styles, PDF export, links with a structure and the right-click menu. All their pictures are new.
0.5.0 19 November 2026
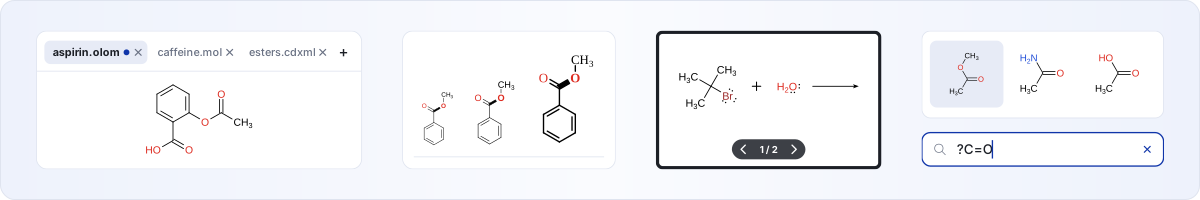
Working with files as in any editor: olomol’s own .olom format, tabs, recent files, PDF, a search bar at the bottom, a presentation mode and a right-click menu. And drawing styles for journals (ACS, RSC or your own).
Added
- Saving: Ctrl+S saves the drawing back into its file without asking (Chrome, Edge), Ctrl+Shift+S is Save as. Drawings are saved in olomol’s own format, .olom; KET files still open and export (for Ketcher, with abbreviations as its groups and the stereo of Haworth and Fischer projections as wedges), but other programs’ files are never written over.
- Tabs: several drawings at once, each with its own undo and file; all of them are back the next time you open olomol. Duplicate drawing (right-click a tab) makes a copy, say of an exercise for its solution. A drawing open in one olomol window stays in it, so two windows never write over each other.
- Opening: drop files onto the window, Open lists recent files, and the installed app opens files from the file manager.
- Search in the bottom bar (/): structures by name, SMILES or InChI, elements, templates, and olomol’s tools and actions.
- Export → PDF / Print…: the drawing alone on a page, as a vector PDF.
- Earlier versions of a drawing: click Autosaved in the top bar, or from the Open menu.
- Show (the button on the right of the top bar): the open drawings full screen, without the bars; → and ← (or a clicker) step through them, from an exercise to its solution, Esc goes back.
- Right-click on the canvas: cut, copy, paste, delete, Redraw, Turn in 3D, flips, a bond horizontal or vertical and more, depending on what you click.
- SVG and PNG pictures exported from olomol carry the drawing: they open again as the drawing (Ctrl+O or dropped on the window).
- Links with a structure:
olomol.org/#smiles=CCO(or#name=caffeine) opens it drawn, several in tabs of their own; a link can be pasted into the search bar too. Chatbots learn how to write them fromllms.txt. - Styles: Settings → Style has RSC, Slides and Custom… (bond length, lines, label size and font in points) beside Screen and ACS. A drawing keeps its style in its file; Use for new drawings sets the style new drawings start in. Circled charges (⊕ ⊖) and lone pairs as bars belong to the style too.
- Bold bonds and bold atom labels (in the selection’s panel), and a text’s own font and size (Aa and pt in the bar over the text).
- While a box is dragged, what it will select is lit before you let go.
- A triple click selects a whole row of a reaction scheme: its molecules, + signs and arrows with what is on them.
- Groups: Ctrl+G groups the selection, groups into larger groups too; Ctrl+click selects a group (again: the group round it), while a plain click still selects atoms and molecules; Ctrl+Shift+G takes a group apart. The same from the Objects panel at the bottom left (a list of the drawing’s things and groups, where groups can be named and things dragged between them) and the right-click menu; Align moves groups as wholes, and SVG pictures keep them as groups.
- A carbocation without its C written: Carbons → line draws a charged carbon as only the + at the corner (− for a carbanion), All C as C+.
- Three new examples in the help, under Files and tabs: an exercise and its solution in tabs, saving and earlier versions, a journal’s style.
Changed
- The top bar’s buttons are in groups. In a narrow window, what doesn’t fit goes under ⋯ at the end of the bar.
- The hint above the bottom bar is one line; a tool button’s tooltip says all the tool does.
- With two drawings or more, the name is on the tabs only; double-click a tab to rename it. New tabs go at the end.
- Keep together is now One compound (and Keep apart Separate), so as not to be taken for grouping.
- Exported PNG pictures carry their resolution, so Word and PowerPoint place them at the size the drawing’s style gives, not four times larger; Copy as picture also puts an SVG on the clipboard.
0.4.5 19 November 2026
Other programs' files: dative and hydrogen bonds, polymers with n, hydrogens set by hand and text formatting are kept. Export says beforehand what a format can't hold.
New
- The Export menu says under each format what of the drawing it can't hold (curved arrows, texts, brackets, colours…), and the message after the export says it again.
Fixed
- Other programs' files (Molfile, RXN, SMILES) hold one whole reaction: a molecule off the arrow's line is kept, resonance, dipole, equals and crossed arrows are no reaction, and a scheme of several steps is written as its molecules (Export says so).
- A reaction SMILES pasted keeps its stereo; olomol writes and reads SDF and RXN itself (a nitrogen with three unpaired electrons stays N).
- ChemDraw captions with a superscript (H⁺, NAD⁺) are no longer lost when opened; InChI of a drawing with an R says why it can't be made, instead of an empty file.
- Molfiles keep dative and hydrogen bonds; read back, they are no longer ordinary bonds (cisplatin with NH₂).
- Brackets with n (a polymer's repeating unit) from ChemDraw, Ketcher and molfiles are kept, and written to molfiles, SDF and KET so that other programs know them.
- Hydrogens set by hand are no longer lost when reading SDF, RXN and Ketcher's files (a carbene CH₂ stays CH₂).
- Texts from ChemDraw and Ketcher keep their sub- and superscripts, bold and italic (m², cm⁻¹, t-BuOK).
- A lone CH₄, NH₃ or H₂O from ChemDraw is an atom, not a label; D, T and R# in a V3000 molfile are read as in V2000.
- A change made just before the page is closed or reloaded is kept; where the browser keeps nothing (a private window), it asks before closing.
- Ctrl+V with a picture on the clipboard says there is nothing to paste, instead of pasting the last copy again.
- Draw out and Abbreviate keep curved arrows on a label and on its bond (OTs leaving in an SN2) and the text on it.
- A label that would run into a nearby atom is written the other way round: HO beside a drawn H, HOH₂C above an OH.
- R/S marks stay off bonds and labels, and out of a ring where they would read as another atom's (glucose as a chair).
- Redraw keeps the side of a drawn H even where its carbon is no stereocentre (a chair's axial H, anti to the Br in an E2): a wedge or a hashed wedge.
- The two fishhooks out of one bond (homolysis) are a mirror pair, on the open side of the bond.
0.4.4 14 November 2026
The preview of a pasted arrow or + in the right place, curved arrows off triple bonds' lines, and one thing done from different places does the same: buttons, keys and the panel, aligning with brackets, turning brackets and texts.
Fixed
- The see-through preview of a lone arrow or + being pasted stands where the click puts it, not off to the left.
- Curved arrows keep clear of the outer lines of double and triple bonds too, and so does a head pointing at an atom (it lay on a triple bond's line).
- Align reaction scheme keeps brackets round resonance forms together and lines up the reaction inside them, also with only the bracket selected: the resonance arrow no longer ends up in front of the bracket, and no wide gap is left after the first molecule.
- The brackets button on the left (and the K key) puts brackets round the selected molecules, as the selection panel's Brackets does.
- Clicking the δ− or δ+ that is on in the selection panel takes it off, as with the δ tool (before, a δ could only be removed by way of the charge).
- Dropping an atom onto another no longer empties the page when a curved arrow starts on one of its bonds; arrows, orbitals and brackets go over to the atom it joins.
- Exporting, copying as a picture and printing only the selection work with a curved arrow selected without its molecules.
- Hide drawn H keeps an H whose bond a curved arrow starts from (an E2).
- Draw out and Abbreviate keep a molecule's place on a reaction arrow, the number under it and its brackets.
- A double click on one resonance form in a bracket no longer selects the bracket round both; the lasso selects brackets as a box does.
- Align works on things inside a bracket, and a bracket in a bracket aligns whichever was drawn first.
- Align reaction scheme keeps a + between molecules joined by a curved arrow (HO⁻ + CH₃Br in an SN2).
- Carbons → line takes the CH₃ off the chain ends too when Settings has All C.
- A clicked bond takes 1 to a wedge, as a bond under the pointer does; 1, 2, 3 make a hydrogen, dative or partial bond a plain one.
- The charge, unpaired electron and bond buttons on the left, the elements in the bottom bar, the element keys and . act on the selected part of a molecule, as their keys and the panel do.
- A charge from the tool or a key takes an old δ off (it came back) and goes up to ±8 as in the panel; a δ on a charged atom replaces the charge.
- Del with the pointer on a side arc deletes that arc, not the whole reaction arrow; a selected side arc stays selected.
- The arrow keys move a compound number or a reagent off its place, as a drag does (nothing moved before).
- In the panel: Lone pairs lights up for a whole molecule, an aromatic bond shows no order and no 2nd line to move, a wedge on a dashed bond makes a plain wedge, a bold text can be edited in its field, a typed length of an arrow with a side arc stays, Other way goes round all the ways.
- Brackets drawn as a box stay upright round what is in them when it is turned, turned over a bond, made horizontal or vertical, or redrawn; texts turn about their middle, and Flip leaves them where they are.
- A flipped Fischer or Haworth projection says it is the mirror image, and an HO written on the left becomes OH on the right.
- A copied bracket made round a molecule includes the atoms drawn on later; a Newman projection's bond seen end on stays single with the bond tool too.
- A charge in a ring's middle is circled when charges are; Fit counts curved arrows in.
0.4.3 13 November 2026
Reliable R/S for chairs and Haworth projections, keys for a whole selection, abbreviations for parts of a selection, and an about page with common questions.
Changed
- With a selection, + and − change the charge of all its atoms, 1 2 3 the order of all its bonds.
- Abbreviate also shortens groups inside a selection that has no abbreviation as a whole (CH2N(CH3)2 → CH2NMe2), and a CH3 selected on its own to Me; Draw out and Abbreviate keep colour and highlight.
- Reagents and conditions beside a turned or upright arrow stand one under the other, close to the arrow, as on a horizontal one.
- The about page answers common questions and, like What’s new, has a page in each language. olomol is cited by one DOI for all versions.
Fixed
- Chairs: a ring carbon with two groups on a tip of the chair, and an axial group with its H drawn, were read with the wrong R/S (cis and trans swapped), in SMILES, InChI and files too.
- Haworth projections with both groups of a carbon drawn (OH and H) were read without R/S, and drawn a little askew with the wrong one. Now the ring’s middle carbons are read, and none wrongly.
- Ring flip works on a flat ring with a wedge on a ring bond (tramadol), and with a carbon with two groups on a tip.
- Align reaction scheme keeps a molecule in the row when a curved arrow goes from it to a reagent on an arrow.
- The + or − in front of a reagent on an arrow stands level with it and in front of its count.
- : with a selection shows its lone pairs, as the button’s tooltip says.
- The tips on the example drawings stand beside the pointer.
- A lone radical atom from a SMILES (
[Cl]) has its unpaired electron. - Fit everything shows a drawing with a long note of several lines, instead of zooming out until nothing can be read.
- A text on an arrow or under a molecule, double-clicked, is edited where it is drawn, not a little to the side; the text tools over it no longer run under the panel on the right.
0.4.2 10 November 2026
A citation with its DOI in the About window, files from ChemDraw and reactions from RXN files read properly, a new address for problems and ideas, and the libraries’ licences complete.
Added
- How to cite in the About olomol window: the citation of the version you use, with its DOI on Zenodo, to copy as text, BibTeX, RIS or CSL JSON or to download as a .bib, .ris or .json file for a reference manager.
Fixed
- Files from ChemDraw (CDXML, CDX) all open, also those whose stereochemistry doesn’t add up. Abbreviations (OMe, OTBS, Et) stay abbreviations (in Ketcher’s files too), labels ChemDraw itself couldn’t read stay as written (SiR3), an atom with a label has the hydrogens its label says (a lone C stays C), and ChemDraw’s attachment point (the squiggle across a bond’s end) becomes an R.
- Reactions from RXN files and reaction SMILES: the plus and the arrow no longer run into labels, the reagents go onto the arrow, and the arrow is only as long as they need.
- R groups (R1), atom lists ([C,N]) and double bonds with their E/Z left open (in KET from Ketcher too) are read as such, no longer as carbons.
- Export through Indigo (CDXML, RXN, SDF): texts are kept, and a drawing with a group label or with wedges that contradict each other now exports.
Changed
- Problems and ideas now go to bugs@olomol.org: the bug button’s e-mail, the About window and the manuals all use it.
- The licences of every library olomol is built on are complete, including those built into RDKit and Indigo (InChI, FreeType, zlib and others). The About window links to them.
- The About olomol window is shorter and easier to read, in parts; the keys and the mouse are in the Keyboard shortcuts window, which it links to.
0.4.1 9 November 2026
Fixes after the 0.4.0 release, Templates in a list, charged groups as abbreviations, a colour for an orbital's other phase, and the MIT License.
Fixed
- Redraw: a molecule with a double bond whose E or Z the drawing leaves open (a substituent in line with the double bond, as when a group is dragged far off) got no drawings, and the window said every engine gave the current drawing. The drawings now keep it open with a wavy bond beside the double bond, as a pasted SMILES has. When every drawing would change the molecule, the window says so.
- A dashed overlap line to an s orbital (a hydrogen in hyperconjugation) meets it on the side facing the other orbital; before, it always started on the right of the atom, even when it ran to the left.
- A dashed overlap line stays between the lobes you dragged it between, say from a C–H bond to the lower lobe of a p orbital; before, it always joined the two lobes nearest each other.
- Draw out: a group like CHO, OMe or Et lies on the 30° grid; before, an aldehyde's C=O was 2° askew.
Changed
- Licence: olomol is under the MIT License, to use, share and change. The About window links to its text.
- Orbitals: the lobe of the other phase can have a colour too. The paint tool colours the lobe you click; in the panel, a toggle above the colours, shaded or other phase, chooses which lobe they paint.
- Templates: the groups are a list on the left with the search above it, and the large ones are split into subgroups (rings: aromatic, heteroaromatic…). Conformations and projections are one group, by how they are drawn: chairs and boats, Newman, Haworth and Fischer projections. Adamantane is drawn flat, and the adamantane cage has its bond behind broken where a bond in front crosses it.
- Charged groups as abbreviations: SO3−, CO2− (COO−), OPO32−, OPO3H− and PO32− stand for a sulfonate, carboxylate or phosphate with the charge where it belongs. Abbreviate writes a selected sulfonate or carboxylate as one, and Draw out brings it back.
0.4.0 8 November 2026
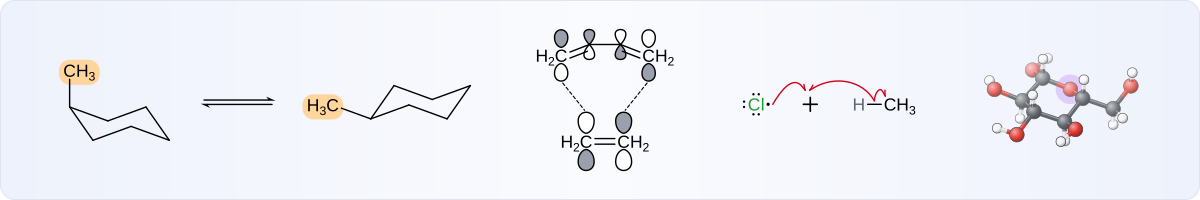
Conformations (the ring flip, Newman projections as molecules, chairs with stereochemistry), orbitals and arrows for radical mechanisms, a stick model in 3D, complexes worked out by physics, and a tidier window.
New
- Ring flip (a button in the top bar): click a chair and it becomes the other chair, axial groups equatorial and the other way round; each group stays up or down, so cis/trans and R/S stay. The flip plays through a boat (Settings → Animations turns it off); with Shift the other chair is drawn beside it with an equilibrium arrow. Cyclohexane in a Newman projection flips too. A flat six-membered ring (a Haworth projection too) clicked becomes the templates' chair (a sugar ⁴C₁): its groups go axial or equatorial by their wedges, those whose side isn't drawn go equatorial, with a wavy bond on a stereocentre. Turn a chair, next to it, turns a chair a third of the way round (Shift: the other way), each group staying axial or equatorial.
- Newman projections are molecules: the front carbon is the point in the middle, the back one the circle, and the groups round them are ordinary atoms. Curved arrows, charges and R/S work on them, so an E2 can be drawn right in the projection; the panel changes the groups and the dihedral angle. Templates → Conformations has cyclohexane in a Newman projection.
- Chairs carry stereochemistry: a chair, from the templates or drawn by hand, is read as the molecule it shows, so SMILES, Info, PubChem and files get the stereoisomer drawn; a chair recognised turns green-teal under the pointer. Turn in 3D keeps the conformation drawn: an axial group stays axial, a Newman projection keeps its dihedral angle, a diene drawn s-cis stays s-cis.
- Orbitals on atoms for mechanisms: p, hybrid and s orbitals, with a phase and lobes sized by the coefficients. A curved arrow can start or end on a lobe, and a dashed line shows an overlap, such as the HOMO and LUMO of a Diels–Alder reaction. Help's examples show them.
- Arrows: while you draw a fishhook, or a curved arrow with Shift, faint lines lead to the atoms the electrons could make a new bond with, and an arrow dropped on the dot in the middle ends between them, so hydrogen abstraction and two radicals combining look as they do in textbooks. An equilibrium arrow can lie to one side (Favours in the panel), a side arc on it meets the harpoon on its side and goes its way (above, forward; below, back), and the reaction arrows have a crossed arrow (the reaction does not go) and an equals sign (the same compound drawn another way).
- Lewis symbols of atoms: up to four unpaired electrons, each on a side of its own as a Lewis symbol fills them (·N̈·, ·Ċ·), and a singlet's two side by side; Unpaired in the panel. An unpaired electron sits to the right of its atom where there is room, and a fishhook from the atom starts at it.
- Stick model in Turn in 3D (S): the model as balls and sticks, with every hydrogen, in the drawing's colours and highlights; shorthands (Ph, OBz) drawn out, G shows each as one ball, H the hydrogens it adds small. It can be saved as a picture.
- Metal complexes in 3D are worked out as a whole by a force field, so porphyrins are flat and chelates sit as they should, in Turn in 3D and in XYZ files. Octahedral complexes with chelates get Λ or Δ beside the metal (R/S, E/Z, Λ/Δ).
Changed
- The top right has the bug, for reporting a problem in one click, and Settings, with the theme and debug mode in it. Help's header has the manuals, What's new, the keyboard shortcuts and About; the bottom left corner the version with its date, What's new and About.
- Drawing: a chain grows across the page, and a click with Shift puts a new bond on the other side (the chain tool's zigzag too). The charges are one button with +, −, δ+ and δ−; a tool's variants open when the pointer rests on its button; D goes through the dative, hydrogen and partial bond. Next in the panel moves a double bond's second line on.
- The Turn in 3D window is larger and stays still as you switch views. The dashed aromatic style draws a dashed circle.
- An element symbol in the panel is taken as written or in small letters: CO is no longer cobalt.
- Some examples in Help are redrawn, the radical chlorination of methane has its termination step, and two new ones are on chairs: a flat ring becoming a chair (menthol) and E2 on a chair.
- A molfile, SDF or SMILES file opened goes into the drawing you have, placed like a template, as an XYZ file does; a KET file opens as the drawing. Export no longer offers RXN and CDXML until they are tried in the programs that read them.
- Redraw lines every drawing of a whole molecule up with the page: turned by up to 15° so that most bonds lie at multiples of 30° (a view of a 3D model keeps its own), and a molecule pasted as SMILES is lined up the same way, so the "straightened" variants of RDKit and CoordGen are gone. Drawings from a 3D model are offered only for cages and bridged rings, metal complexes, molecules with an expanded octet and SN2 transition states; Turn in 3D shows any molecule in 3D. Redraw also offers more good drawings: some it used to drop for a wedge that read the wrong way. The drawings in its window no longer change places when a slower one arrives: it is added after the others.
- When the drawing of a structure from a SMILES, from PubChem or from an XYZ file has atoms on top of each other or crossing bonds, the tidiest of Redraw's quick layouts is taken instead.
Fixed
- Metal complexes drawn from their 3D model (Redraw, Turn in 3D) have long bonds to the metal again (1.8 bond lengths), so the labels round the metal have room; a chelate is made larger to reach (at most 1.6×). An XYZ file gets the real geometry. Turn in 3D again draws bonds seen nearly end on long enough to read, as Redraw does (but for a transition state, whose partial bonds shorten as it turns).
- Names typed under molecules stay with them, in a grid and in Align reaction scheme.
- The aromatic circle lies in the ring's plane, in a ring turned away or drawn out of shape too.
- Hydrogens drawn by hand no longer give wrong bond lengths in 3D.
- A shorthand on a coordination bond (PPh₃) counts in full in files, SMILES, InChI and Info.
- An abbreviation such as Ph can no longer get a second bond; one in an older drawing is ringed red.
- Flip over a bond turned the stereocentre at the end of a wedge the other way: maltose became cellobiose.
- A pasted SMILES sometimes changed the molecule: artemisinin lost the stereocentre at its peroxide bridge, and some macrolides got a diene in the ring the wrong way round. Now another drawing is taken then. A double bond whose E or Z the SMILES leaves open gets a wavy bond beside it (before, the drawing decided E or Z), and Redraw keeps it. A double bond marked "either" in a molfile comes in the same way.
- A metal complex's XYZ file had a broken geometry; Redraw of a chair offers every drawing; in 3D an axial hydrogen stays axial.
- In Turn in 3D a transition state turns as a thing in space, and sulfuric acid and the like have all their hydrogens in the stick model. Draw the H on a chair puts each hydrogen axial or equatorial, as the templates do.
- A cyclohexene drawn in a chair's shape (an E2 product) keeps its R/S in files, SMILES, Info and Redraw.
- On a Czech keyboard + over an atom raises its charge (it is on the 1 key). The keyboard shortcuts (?) list Turn in 3D's keys and those for writing text. Esc closes a tool's variants and leaves the tool; a click outside Help closes it instead of drawing under it. The Formula button writes e- as e⁻.
0.3.0 2 November 2026
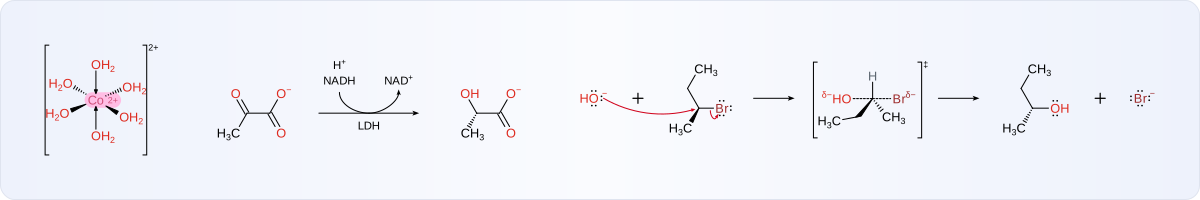
Turning molecules in 3D, redrawing a selected part of a molecule, a new Help with examples to point at, searching templates by a part, easier texts, wedged coordination bonds and side arcs on reaction arrows.
New
- Turn in 3D (the cube next to Redraw): turn the selected molecule with the mouse or the arrow keys and see it exactly as it will be drawn, its back paler. Use this view draws it so, with wedges that keep its R/S and E/Z. The 3D model is checked against the drawn stereochemistry. It warns when a view would lose R/S or E/Z, and the view can still be used. Complexes have each ligand in its proper shape and chelating ligands (EDTA) wrapped round the metal; where two bonds cross, the one behind is broken. Redraw also draws an SN2 transition state as textbooks do (a trigonal bipyramid).
- Redraw (formerly Layout): with only part of a molecule selected, it offers to redraw just that part, the rest staying exactly where it is. These drawings have a section of their own above those of whole molecules. Drawings are ranked by crossings, overlaps and uneven bonds and angles: the best come first, the worse ones wait behind Show all drawings. 3D drawings of large molecules (taxol) are worked out in the background, so the page doesn't freeze, and side by side, so they are ready sooner.
- Help (the round ? button at the bottom right) opens at once: twenty-five finished drawings with pictures, from mechanisms through sugars and complexes to texts and export, to pick from by their tags. Point at any part to see how it is drawn; its button lights up. Under a drawing is what can't be pointed at (Redraw, Templates, Align). Any example opens as your own drawing. The window's header has the quick start, the manual, what's new, the keyboard shortcuts and reporting a problem. Four examples take turns on the empty canvas.
- Searching templates by a part: ?oh, ?Ts or ?ester lists the templates with that group, ?ala finds alanine by its code; capitals don't matter. Elements, abbreviations and parts written as SMILES work too (?S, ?boc, ?C=O). Amino acids have their codes on their cards, and without the question mark a code finds them first (Phe, F).
- Texts: the window over a text has Formula, Plain, subscript and superscript, bold and italic (also for part of a text), line alignment and a characters menu (Greek letters, arrows, ° ± ‡) in one row. Formula also handles H₂[SiF₆]²⁻ and CO_3^2-. A copied text pastes into other programs as plain words (a DOI into the browser, say). A multi-line text on a molecule or an arrow is centred by itself.
- Wedged coordination bonds, plain or hashed, for complexes in space (Λ and Δ), without a warning about the number of bonds. The 3D views keep the arrangement as drawn. Settings can draw coordination bonds as lines instead of arrows.
- Side arcs on reaction arrows (NAD⁺ → NADH): with the reaction arrow tool, drag from an arrow's middle outwards, or from the canvas to it. A text or a molecule goes at the arc's end, and the arrow grows to fit. The arrow's panel turns it round or flips its arcs to the other side.
- PubChem: when a drawing leaves part of its stereochemistry open, or PubChem doesn't have exactly that compound (other stereochemistry, isotopes), PubChem page shows a page of cards of similar compounds (picture, name, CID) and says what differs. The cards have names instead of database codes.
- An autosave mark beside the drawing's name: Autosaved means the drawing is in this browser and will be here when you come back. It warns when the browser can't autosave (a private window, full storage) and when another tab with olomol has autosaved its drawing since: next time the one changed last comes back.
- Isotopes: type 13C, 18O, D or T into the Element field; the formula and masses count them.
- The quick start and the manual in English.
- An about page (linked from the window about the app): what olomol is and does, with four short videos.
Changed
- XYZ files are worked out in the background: keep drawing, and when one is ready, save it with Save XYZ file. Its InChIKey is suggested as the name, and the panel warns about stereocentres with no R/S drawn.
- The 3D model for Turn in 3D, Redraw and XYZ files now comes from RDKit: one try at taxol takes about 2 seconds instead of 8. The result is the same in every browser, though large molecules can differ a little.
- Abbreviations and group labels moved from the periodic table to Templates (the Abbreviations tab). A functional group's card has a chip with its abbreviation (Ph on Phenyl) that puts the group in as one atom. Shift+T also opens Templates.
- Rings and functional groups: dragged from an atom, they point at the pointer in 15° steps. A ring hung on an atom gets atoms of its own, so three clicks of benzene on the P of P=O give Ph₃P=O with no rings fused. With Shift, a group's first atom becomes the clicked atom (COOH on pentane's end carbon gives pentanoic acid) and a dragged ring goes through the atom (spiro).
- Abbreviate handles several groups and whole molecules: the phenyls of PPh₃ become three Ph, and a molecule such as PPh₃, NEt₃, Et₂O or AcOH becomes one label. A label typed that way counts as its molecule in formulas, Info, PubChem and files. In labels, letters that are atoms take their element's colour (the P of PPh₃, the O of OBz).
- The unpaired electron and the fishhook arrow are now variants of the lone pair and curved arrow buttons (the triangle in the button's corner), and a fishhook has a real barb.
- Horizontal / vertical is now a tool in the top bar: click a bond and the selection turns if the bond is in it, else the whole molecule; another click turns it a quarter further. With one bond selected, its molecule turns at once.
- Key 1 over a single bond goes through wedge, hashed wedge, both from the other end, and plain; Shift+1 goes back. Esc, with nothing left to cancel, switches between box and lasso selection. R pressed again gives the next ring.
- Shift+double-click adds a molecule to the selection. Show everything with a selection zooms until it fills the window.
- Lone pairs sit a little further from their letters, a bond that would run through a charge stops at it, and a charge with no free corner goes straight beside, over or under its symbol. Align and Align reaction scheme go by atoms and their symbols, so Cl⁻ and H₂O sit level with Cl and O.
- Faster: on a first visit over a school line, the first SMILES is drawn in about 2 seconds instead of 4.5; large drawings drag smoothly, and the panel no longer slows down with a large selection.
- Esc in a text keeps it instead of dropping it. A new drawing starts at 100 %. olomol starts in the light theme; dark or the system's is in Settings. Pictures are see-through by default, and a white background is remembered once ticked. The panel's charge has − and + buttons, for any charge.
Removed
- Tilt back: the button is gone. Turn in 3D shows a complex or a molecule in perspective.
Fixed
- Exporting a selection with abbreviations (Ph, Boc) no longer fails, and keys such as Delete and Ctrl+Z no longer change the drawing behind an open window.
- Abbreviate and Draw out keep the wedge of a group on a stereocentre, so the stereochemistry stays (cocaine's OBz, say, which PubChem finds again).
- A click of the chain tool on a bond no longer breaks it. Flipping also swaps the side of a double bond's second line.
- XYZ files: onium cations read back correctly, and a lone ion no longer stops an export.
- SMILES files: dots stay in names, and rings no longer close across a dot.
- A failed engine download is tried again, and error messages are no longer half in English.
- The drawing's default name is no longer translated, a charge or δ no longer sits on a double bond's second line, and brackets keep fitting when an atom in them is deleted.
0.2.2 29 October 2026
Ketcher opens KET files again.
Fixed
- A KET file saved in olomol opens in Ketcher (it used to say it couldn't read the file). Ketcher checks a file against its schema and objected to where the part with what only olomol uses (colours, lone pairs, curved arrows, brackets) was. Now it is where the schema allows it. Files saved earlier open in olomol as before.
0.2.1 29 October 2026
Fixes to XYZ files (3D coordinates), this list of changes, and a filled-in e-mail for reporting a problem.
New
- This list of changes, in the help menu (the round ? button at the bottom right) under What's new.
- The e-mail from the report window (the envelope at the top right) now has a table with the version, browser and system. Only the link to the drawing is too long for an e-mail: Copy the table copies it.
Fixed
- XYZ export keeps the drawn stereochemistry (R/S, E/Z). In crowded ring systems (such as the spiro carbon of hinckdentine A) another diastereomer could come out. Every 3D structure is now checked, and if the right one isn't found, the export stops with a message instead of writing a wrong molecule.
- XYZ export also works for molecules with boron or an acylium cation (C≡O⁺), for which the MMFF94 force field has no atom types. They are written without minimisation; the file's comment line says so.
- XYZ import puts a radical where it belongs: a phenyl radical on the carbon without a hydrogen (it used to load as a CH radical), a benzyl one on the CH₂.
0.2.0 28 October 2026
Anchors for texts and molecules on molecules and on reaction arrows, a multistep arrow and new templates.
New
- Anchors on molecules: every molecule has four places for text (below, above, left, right). Drag a text to it: it snaps there and then moves and aligns with the molecule. A compound number goes below, a count (2 NaOH) on the left. Several texts on one anchor keep the order you put them there in; drag to change it.
- Places on reaction arrows: drag a text (conditions) or a whole molecule (a reagent) above or below an arrow; they move with the arrow, and dragging changes their order. The panel puts + or − before a molecule on an arrow (added, taken away). A double click on an arrow selects it with everything on it.
- Multistep arrow (two arrows one over the other, one object), for "in n steps".
- Templates: adamantane as a cage (besides the flat one).
- With an element chosen, a click on the empty canvas puts down the atom alone (O: water).
- A filled ring can be clicked: a click selects the ring, a double click the whole molecule, dragging moves it.
Changed
- The Compound number tool is gone: a bold text on the anchor below a molecule does the same and can be edited freely.
- Align goes by the anchors: a text on an anchor goes with its molecule, free texts are aligned themselves, so several texts and molecules can be aligned at once. Align reaction scheme makes an arrow as long as what is on it.
- The grid takes molecules row by row from left to right, as you see them.
- Chairs (sugars' too) have their tips level with the top and bottom corners.
- Highlights of different colours: the smaller mark is always on top.
- The arrow kinds in the panel have the same symbols as in the bar.
Fixed
- Typing after Shift+Enter no longer adds a new line with every letter.
0.1.1 28 October 2026
Texts are formatted only where you want it.
Changed
- What you type stays as it is: digits in a DOI or in names no longer become subscripts by themselves.
- You format a formula yourself: select part of a text and click Formula in the window over it (H₂SO₄, Fe³⁺); Plain takes the formatting off. Subscripts show while you type.
- Older KET files are converted when opened, so they look as before.
0.1.0 27 October 2026
The first version: a chemistry drawing program that runs entirely in the browser.
Drawing
- Bonds (single, double, triple, wedges, hashed wedges, wavy, dative and hydrogen bonds), chains, rings of 3 to 8 atoms and benzene, with a see-through preview of what will happen.
- Elements from the bottom bar, the periodic table and keys; abbreviations (Ph, Boc, OMe…) and generic groups (R, Nu, LG).
- Charges (δ+ and δ− too), radicals, lone pairs and hydrogens, which the program counts itself.
- Text with subscripts and superscripts; reaction, equilibrium, resonance and retrosynthesis arrows, a dipole arrow, plus signs.
Mechanisms
- Lone pairs and unpaired electrons; curved arrows and fishhooks tied to atoms and bonds, which find their own way round the drawing.
- Resonance structures, transition states and brackets; aromatic rings as a circle or dashed (aromatize and Kekulé).
Chemistry
- Formula, molar and exact mass, elemental analysis and ions for HRMS.
- R/S and E/Z on chosen atoms and bonds, IUPAC numbering with the reasons, an Info card with SMILES, InChI and a PubChem link.
- A check that edits don't change the molecule; an atom that doesn't fit is marked.
Templates and structures from a name
- Rings, functional groups, compounds, conformations (chairs, Newman projections), projections (Fischer, Haworth), complexes, amino acids, sugars and nucleobases, reagents and solvents.
- A structure from its name, SMILES or InChI (names are looked up in PubChem).
Editing and arranging
- A new layout with several drawings to choose from (from a 3D model too), aligning, spacing evenly, a grid and a reaction scheme in one row, flipping.
- Colours, highlights and ring fills; Keep together for salts.
Files
- Saving as KET (Ketcher opens it too); export to molfile, SMILES, RXN, CDXML and XYZ (3D), SVG and PNG pictures; a picture into Word, PowerPoint and Google Docs with Ctrl+Shift+C.
- A link that carries the whole drawing; the drawing saves itself.
The app
- It runs entirely in the browser, nothing is installed and your drawings stay with you; it works offline and can be installed as an app.
- Czech, English and German; light and dark looks; keyboard shortcuts; a quick start and a manual in PDF; reporting a problem by e-mail with a table of details.